I spent the first six years of my career chasing “impossible” results before I learned that most analytical failures are not mysteries. They are boring, repeatable, and almost always traceable to a consumable, a connection, or a preparation step that nobody wrote down. The instrument gets blamed first because it is expensive and loud. The vial, the septum, the syringe filter, and the torque wrench are where the money actually leaks. What follows is the failure catalog I keep taped inside a lab cabinet, drawn from hundreds of investigations across HPLC, GC, ICP-MS, and wet chemistry benches.
When a sequence fails, the instinct is to open the instrument log. I do the opposite. I ask three questions: what touched the sample, what touched the column, and what changed since the last passing run. In my records, roughly 60 to 70 percent of out-of-specification results trace back to sample preparation or sample handling, not the detector. Another 15 percent come from mobile phase or carrier gas problems. The instrument itself is the culprit less often than the paperwork suggests.
That split matters for how you spend your maintenance budget. A new detector lamp costs more than a year of vials. A column costs more than a decade of septa. Yet the cheap consumables are where I see the most corner-cutting, because they look interchangeable and the failure they cause is delayed and indirect. If you are building a prevention program, start at the bench, not the service contract. The full scope of what a testing lab handles, from method development to release testing, is worth mapping against your own failure log at Analytical Testing at labanalyticaltesting.com.
Failure 1: autosampler vial and septum problems
What goes wrong
The classic symptom is a ghost peak that appears in blanks and grows through a sequence. The cause is usually a septum that has been pierced too many times or a vial that was never designed for the solvent in it. A standard 11 mm crimp or snap septum is rated for a limited number of injections before the puncture channel stops resealing. Once that channel opens, the headspace exchanges with the room, volatile analytes escape, and oxygen and moisture get in. I have watched a 2 percent loss of a volatile analyte per hour become a 15 percent drift over an overnight run.
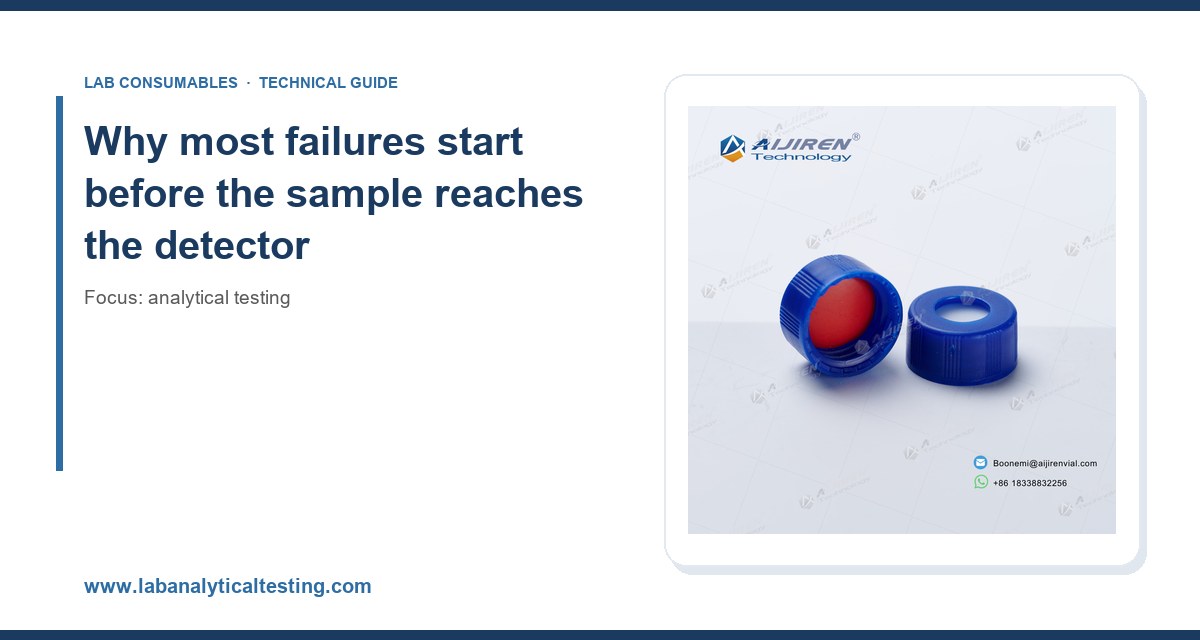
Septum material selection is not cosmetic. Silicone is soft and reseals well but leaches siloxanes that show up as m/z 73, 147, 207, 281 in GC-MS. PTFE-faced silicone gives you a chemically inert sample contact surface and a resealing core. Natural rubber is cheap and terrible for anything acidic. For LC-MS work, a pre-slit silicone/PTFE septum reduces coring and the associated particles that end up on the column inlet.
How to prevent it
- Cap the puncture count. I set a hard limit of 3 to 5 injections per septum position for volatile work and 10 for aqueous LC. Rotate vial positions in the autosampler tray so no single septum takes the whole sequence.
- Match the vial to the solvent. Use amber glass for light-sensitive analytes, and confirm the glass is a low-extractable borosilicate. Clear vials are fine for most work but I default to amber because it removes a variable.
- Check the crimp. A loose crimp leaks; an over-crimped one cracks the neck. Target a crimp that holds the cap firmly with the septum compressed but not deformed. A proper crimping tool with a depth stop beats hand pressure every time.
- Fill to a consistent volume. Inconsistent fill changes headspace and therefore changes partition behavior. I standardize on a fixed fill line, typically 1.0 to 1.5 mL in a 2 mL vial.
For a deeper look at how vial geometry and closure design interact with autosampler mechanics, the resources at Chromatography Vial at chromvials.com and Hplc Vial at hplcvialfactory.com cover the dimensional tolerances that matter. A vial that is 0.2 mm out of spec on neck height can cause needle strikes that you will never see in the data, only in the downtime.
Vial and septum decision checklist
- Volatile analytes (headspace, residual solvents): 20 mL headspace vial, PTFE-faced silicone septum, magnetic or crimp closure rated to 150 °C, single-use.
- LC-MS, trace level: 2 mL amber borosilicate, pre-slit silicone/PTFE septum, low-extractable certified, ≤5 injections per position.
- Routine HPLC, UV detection: 2 mL clear or amber, silicone/PTFE, ≤10 injections per position.
- Ion chromatography: avoid glass entirely; use high-density polyethylene vials to prevent sodium and potassium leaching.
Failure 2: sample filtration that removes more than particles
Syringe filters are the most under-specified consumable in the lab. A 0.45 µm filter is standard for HPLC, but a 0.22 µm filter is what I use for anything headed to a column with a 3 µm or smaller particle bed, because sub-visible particles that pass a 0.45 µm membrane will lodge at the inlet frit. The bigger problem is adsorption. Nylon binds proteins and some basic analytes. PTFE is chemically inert but hydrophobic, so it needs a pre-wet with methanol or acetonitrile before aqueous samples. Cellulose acetate is fine for aqueous but degrades in strong organic solvents.
I have seen recovery of a 1 µg/mL hydrophobic analyte drop to 40 percent after a nylon filter because the compound adsorbed to the membrane. The fix was a PTFE filter with a glass fiber prefilter. The lesson: run a recovery check on any new filter lot before you trust it. Spike a known concentration, filter it, and compare to an unfiltered control. If recovery is below 95 percent, change the membrane.
Filter selection by matrix
- Aqueous, protein-containing: regenerated cellulose or PES, 0.22 µm, low protein binding.
- Organic solvents: PTFE, 0.22 or 0.45 µm, pre-wet with the sample solvent.
- High-particle slurries: glass fiber prefilter stacked ahead of the membrane to prevent premature clogging.
- Trace metals: avoid membranes with metal-containing binders; use PTFE or cellulose.
Failure 3: mobile phase and carrier gas problems
Mobile phase degradation
Buffer precipitation is the silent killer of LC columns. A phosphate buffer at 50 mM in a high-organic mobile phase will salt out at the mixing point if the proportion of acetonitrile climbs too fast. I keep organic content below 70 percent for phosphate systems and switch to ammonium acetate or ammonium formate when I need higher organic. Filter every aqueous buffer through a 0.22 µm membrane and degas it. Dissolved gas causes pump cavitation and baseline noise that looks like detector trouble.
Microbial growth in aqueous buffers is another one. A buffer left at room temperature for three days grows enough biomass to clog a frit. I add a small amount of sodium azide (0.02 percent) for non-MS work, or prepare fresh buffer daily for MS. Never store aqueous buffer in a bottle that previously held high-organic solvent without a thorough rinse.
Carrier gas and GC inlet issues
In GC, an oxygen trap and a moisture trap on the carrier gas line are not optional. Oxygen at the parts-per-million level degrades a polysiloxane stationary phase, and the symptom is a rising bleed baseline that mimics column failure. I replace traps on a schedule tied to gas cylinder count, not to a calendar. A leaking septum or a loose inlet nut lets oxygen in continuously, so check for leaks with a thermal conductivity leak detector at every connection, including the 1/16 inch fittings on the inlet and detector. Torque these fittings to the manufacturer’s spec, typically snug plus a quarter turn, and never overtighten a graphite ferrule, which will crack and leak.
For reference material on column and accessory selection, the Agilent chromatography consumables and columns catalog and the Restek technical resources for chromatography are where I send new analysts to understand phase polarity and bleed characteristics. The Thermo Fisher chromatography and sample handling pages are useful for matching sample prep to detection mode.
Failure 4: column and system contamination
The inlet frit is where particles die. A column that loses 20 percent of its plate count in a week is usually not worn out; it is fouled at the head. Backflushing can recover some of it, but the real fix is upstream: filter samples, use a guard column, and replace the inline filter. I budget a guard column change every 200 to 500 injections depending on matrix, and I log injection count against system suitability results so I can see the trend before it becomes a failure.
Carryover is the other contamination failure. A 0.1 percent carryover from a high-concentration sample into a low-concentration blank will fail a limit-of-detection check. The causes are a dirty injection port liner, a worn needle seat, or a sample loop that is too large for the injection volume. I run a blank after every high standard and set a carryover acceptance limit of ≤0.05 percent of the high standard peak area. When it fails, the liner and needle seat are the first two parts I swap.
Failure 5: calibration and standard preparation drift
Calibration failures rarely come from the instrument. They come from standards that were made wrong. A balance that reads 0.1 mg is not adequate for weighing 1 mg of a reference standard; the relative error is 10 percent. Use a 0.01 mg balance for small weights, or make a concentrated stock and dilute. Check the balance with certified weights at the start of every session, and record the actual weight, not the target.
Standard stability is the other trap. Many organic standards degrade in the autosampler at room temperature over a long sequence. I run a mid-point standard every 10 injections and set an acceptance window of ±5 percent against the initial calibration. If the check standard drifts outside that window, the sequence is invalid, regardless of how good the peaks look. Store standards at -20 °C in amber vials with PTFE-lined caps, and let them equilibrate to room temperature before opening to prevent condensation.
Calibration troubleshooting checklist
- Correlation coefficient below 0.995: check standard preparation, pipette calibration, and injection volume reproducibility.
- Intercept too high: contaminated blank, carryover, or a dirty injection port.
- Nonlinear response at the top: detector saturation or a column overload. Reduce injection volume or dilute.
- Drifting check standard: standard degradation, mobile phase evaporation, or a lamp losing output.
- RSD above 2 percent on replicate injections: autosampler precision, air bubbles in the syringe, or a partially clogged needle.
Failure 6: environmental and instrument drift
Room temperature swings of 5 °C will move a refractive index detector baseline and shift retention times enough to fail system suitability. I keep instrument rooms at 20 to 25 °C with less than ±2 °C variation, and I keep HPLC systems away from HVAC vents and direct sunlight. Humidity matters for balance rooms; keep it between 40 and 60 percent relative humidity to prevent static and moisture uptake on hygroscopic standards.
Detector lamps have finite life. A deuterium lamp rated for 2,000 hours will lose output gradually, and the first symptom is a rising baseline and poorer signal-to-noise at low wavelengths. Log lamp hours and replace at 80 percent of rated life, not at failure. The same logic applies to electron multiplier gain in MS; track the gain and replace before sensitivity falls below method requirements.
Failure 7: method transfer and documentation gaps
I have inherited methods that failed on day one because the original method note omitted the column lot, the mobile phase pH, or the equilibration time. A method is not transferable without those parameters. Write down the column dimensions (4.6 × 150 mm, 5 µm), the pH of the aqueous phase to ±0.1, the equilibration volume (at least 10 column volumes), and the injection volume. When a method moves between labs, run a side-by-side comparison on the same standard before trusting either result.
Regulatory methods add another layer. If your work falls under waste or environmental testing, the method numbers and quality control requirements in the U.S. EPA SW-846 test methods for waste and materials define the blanks, matrix spikes, and duplicate frequency you must run. Skipping those QC samples is itself a failure mode, even when the data looks clean.
Consumable specification reference
The table below is the quick-reference I use when specifying consumables for a new method. Numbers are working targets, not absolutes, and should be verified against your method and instrument manufacturer’s guidance.
- Autosampler vial: 2 mL, 11 mm neck, borosilicate glass, ±0.1 mm dimensional tolerance.
- Septum: silicone/PTFE, pre-slit for LC-MS, puncture limit 3 to 10 depending on application.
- Syringe filter: 0.22 µm for sub-3 µm columns, PTFE for organics, PES or RC for aqueous.
- Guard column: replace every 200 to 500 injections or when back pressure rises 15 percent above baseline.
- Ferrules: graphite for GC, torque to snug plus 1/4 turn; never reuse a deformed ferrule.
- Gas traps: oxygen and moisture, replace per cylinder count, typically every 2 to 4 cylinders.
- Buffer: filter 0.22 µm, store at 4 °C, replace within 3 to 5 days for aqueous systems.
Consumable sourcing deserves the same scrutiny as instrument sourcing. A supplier that cannot provide lot-level certificates of analysis, dimensional data, and extractables information is a risk you are accepting on every injection. I have audited suppliers through Aijiren at aijirentechinc.com and found that the difference between a good and a bad vial lot shows up months later as unexplained drift, long after the invoice is paid.
A practical prevention program
Prevention is cheaper than investigation. The program I run has four parts. First, a consumable change log tied to injection count, not to memory. Second, a daily system suitability check with defined acceptance criteria (RSD ≤2 percent, tailing factor 0.9 to 1.5, resolution ≥1.5). Third, a monthly review of out-of-specification events categorized by root cause so you can see which failure mode is eating your time. Fourth, a supplier qualification file for every critical consumable, including vial dimensions, septum material, and filter membrane composition.
When a failure does happen, resist the urge to replace the most expensive part first. Work backward from the sample. Check the vial, the septum, the filter, the mobile phase, and the fittings before you touch the column or call service. In my experience, that sequence resolves the majority of failures within an hour and saves a service call.
The takeaway is straightforward: analytical failures are usually consumable and preparation failures wearing an instrument costume. Specify your vials and septa with real dimensional tolerances and puncture limits, filter every sample and buffer to 0.22 µm, control your gas and mobile phase chemistry, and log your changes against injection count. Do those four things and the “impossible” results become rare enough that you can spend your time on method development instead of firefighting.
References & further reading
Related resources from our lab consumables network:
- Analytical Testing at labanalyticaltesting.com
- Aijiren at aijirentechinc.com
- Chromatography Vial at chromvials.com
- Hplc Vial at hplcvialfactory.com
External references and standards: